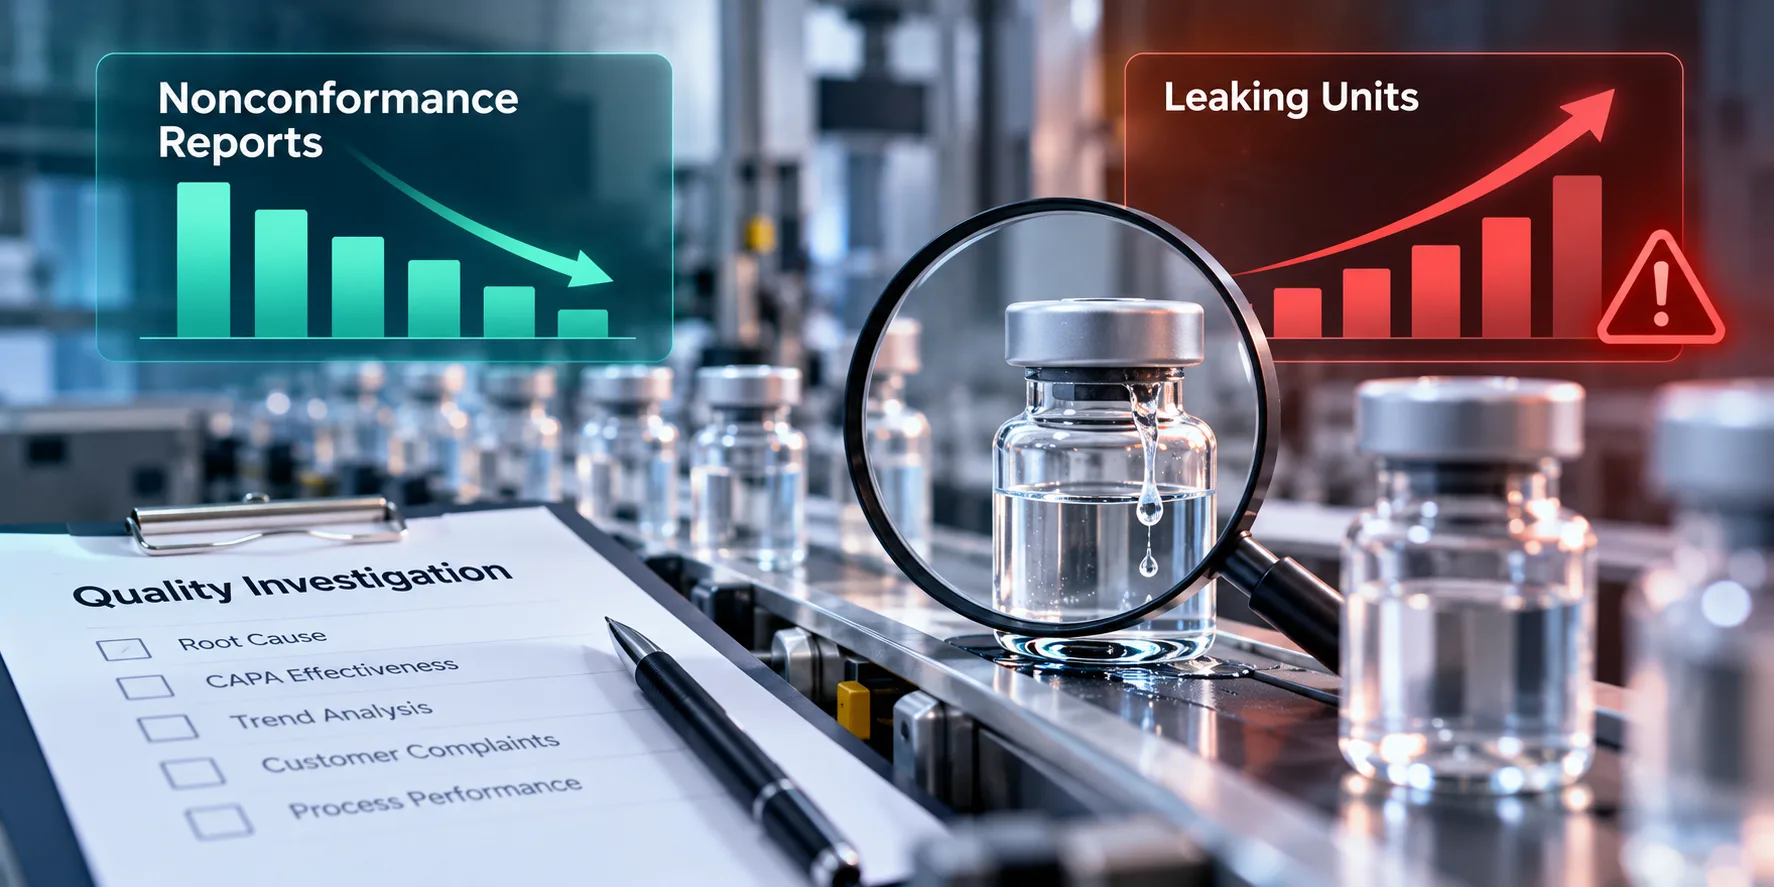
A pharmaceutical manufacturing site reports fewer nonconformances than the previous year. On the quality dashboard, the number looks encouraging. But during the same period, product complaints and manufacturing defects are increasing.
Which indicator should quality leadership trust?
This situation is not hypothetical. A recent FDA warning letter highlights how conflicting quality metrics can expose weaknesses in pharmaceutical investigations and corrective actions.
What FDA Found During a Pharmaceutical Inspection
On September 29, 2026, the U.S. Food and Drug Administration issued a warning letter to Baxter International Inc. following an inspection of its Round Lake, Illinois, manufacturing facility.
Among several significant CGMP observations, FDA identified deficiencies involving container-closure integrity, leaking injectable products, and inadequate investigations.
One observation was particularly revealing.
FDA reported that the rate of leaking units had increased consistently from 2023 through 2026, while the number of nonconformance reports (NCRs) opened had decreased.
The agency also questioned whether previous corrective and preventive actions had adequately addressed recurring failures.
This discrepancy raises an important issue in pharmaceutical quality management: a reduction in reported quality events does not necessarily indicate a reduction in actual product quality problems.
Why Quality Metrics Need Context
Pharmaceutical manufacturers monitor several indicators to evaluate process performance and quality-system effectiveness.
These may include deviation frequency, batch rejection rates, product complaints, environmental monitoring excursions, investigation timelines, and CAPA effectiveness.
Each indicator offers useful information, but no single measure provides a complete picture.
Consider a facility where the number of deviations falls by 20%, while customer complaints involving the same product defect increase.
Several explanations are possible. Manufacturing performance may have changed, detection controls may be inadequate, reporting practices may differ, or investigations may not be capturing the full extent of recurring issues.
The numbers alone cannot establish the cause.
Quality teams need to examine how the indicators relate to one another, whether reporting criteria have changed, and whether trends are being measured against comparable production volumes.
A lower deviation count is useful information. It should not automatically be interpreted as evidence of better quality.
The Role of Investigations and CAPA Effectiveness
Under 21 CFR 211.192, pharmaceutical manufacturers must thoroughly investigate unexplained discrepancies and failures to meet specifications.
An investigation needs to establish what happened, identify scientifically justified causes, assess the scope and potential product impact, and determine appropriate corrective actions.
The work does not necessarily end when the CAPA activities have been completed.
Quality teams must also assess whether the actions addressed the identified causes and whether the problem has recurred.
For example, replacing a defective component may resolve an immediate equipment issue. But if similar failures continue across multiple batches, the original investigation and CAPA deserve further examination.
This is especially important when complaints, production records, or inspection results suggest that problems have persisted despite previous corrective actions.
The ISPE Advancing Pharmaceutical Quality guidance on CAPA systems addresses effectiveness, metrics, management oversight, and governance as important elements of CAPA maturity.
What Should Quality Leaders Review?
When quality indicators appear inconsistent, a useful review extends beyond the number of events recorded during the month.
Quality teams should examine:
- Defect trends: Are particular defects becoming more frequent relative to production volume?
- Complaint patterns: Are customers reporting problems similar to those already observed internally?
- Investigation history: Have related events been investigated separately despite possible common causes?
- CAPA effectiveness: Did previously implemented actions reduce or eliminate the targeted problem?
- Reporting practices: Have event classification, escalation criteria, or documentation practices changed?
- Management review: Are significant adverse trends receiving appropriate attention and resources?
Reviewing these relationships can help identify whether a declining event count reflects genuine improvement or warrants additional investigation.
How Digital Quality Systems Support Better Oversight
Electronic quality management systems can help organizations connect deviations, complaints, investigations, CAPAs, change controls, and related quality records.
Several established software providers address these processes.
MasterControl provides quality event management and CAPA workflows, including reporting and follow-up activities.
Veeva QMS supports quality processes such as deviations, complaints, investigations, CAPA, and change control across life sciences organizations.
epiq QMS by Caliber connects quality processes with document and training management, supporting traceable workflows across related quality activities.
These platforms can help quality teams organize records, follow investigations, and review information across processes.
However, software cannot independently determine whether a root cause is scientifically justified or whether corrective actions have been effective. Those assessments require qualified personnel, appropriate evidence, and responsible quality oversight.
What This Means for Pharmaceutical Quality Management
The Baxter warning letter provides a specific example of why quality performance should not be evaluated using isolated numbers.
A facility may close investigations on time, complete CAPA tasks, and report fewer nonconformances while significant product-quality concerns remain unresolved.
Quality leadership needs to understand the relationship between reported events and the actual performance of manufacturing processes.
That requires reliable reporting, meaningful trend analysis, thorough investigations, and documented assessments of corrective action effectiveness.
The objective is not to increase or decrease the number of deviations reported.
It is to identify quality problems accurately, investigate them appropriately, and establish whether the actions taken have worked.
References
- FDA Warning Letter — Baxter International Inc., September 29, 2026
- 21 CFR 211.192 — Production Record Review
- FDA/ICH Q10 — Pharmaceutical Quality System
- ISPE — Advancing Pharmaceutical Quality: CAPA System
- MasterControl — Quality Event Management
- Veeva — Quality Management System
- Caliber Technologies — epiq QMS